Table of Contents
Why Pharmaceutical Manufacturers Fail Regulatory Due Diligence Before Acquisition


Pharmaceutical acquisitions are complex, high-stakes transactions. Financial modelling, commercial valuation, and market positioning typically receive the most intensive scrutiny. Yet the most consequential risks in many transactions are neither financial nor commercial — they are regulatory.
Regulatory due diligence failures in Pharmaceutical M&A are not rare. They surface repeatedly after transaction close, in the form of manufacturing site warning letters, Import alerts, product registration disputes, or post-acquisition compliance remediation costs that erode deal value significantly. In the most serious cases, they result in product withdrawals, regulatory-mandated shutdowns, or reputational damage affecting the entire acquiring organisation.
The most experienced Pharmaceutical investors treat regulatory due diligence not as a compliance checkbox, but as a fundamental component of investment thesis validation.
1. Why Regulatory Due Diligence Is Distinct from a GMP Audit
A GMP audit evaluates compliance status at a point in time. Investor-grade regulatory due diligence goes significantly further:
- Complete audit of all product registrations, manufacturing licences, and Import/Export authorisations across every market
- Full inspection history — CDSCO, USFDA, EMA, TGA — including correspondence and whether commitments were fulfilled
- QMS maturity assessment as a sustained operational discipline, not just a documented framework
- Data integrity posture across all GMP-critical systems
- Regulatory liability exposure from pending variations, renewals, or disputed dossiers
- Regulatory risk concentration — over-dependence on a single product, site, or market
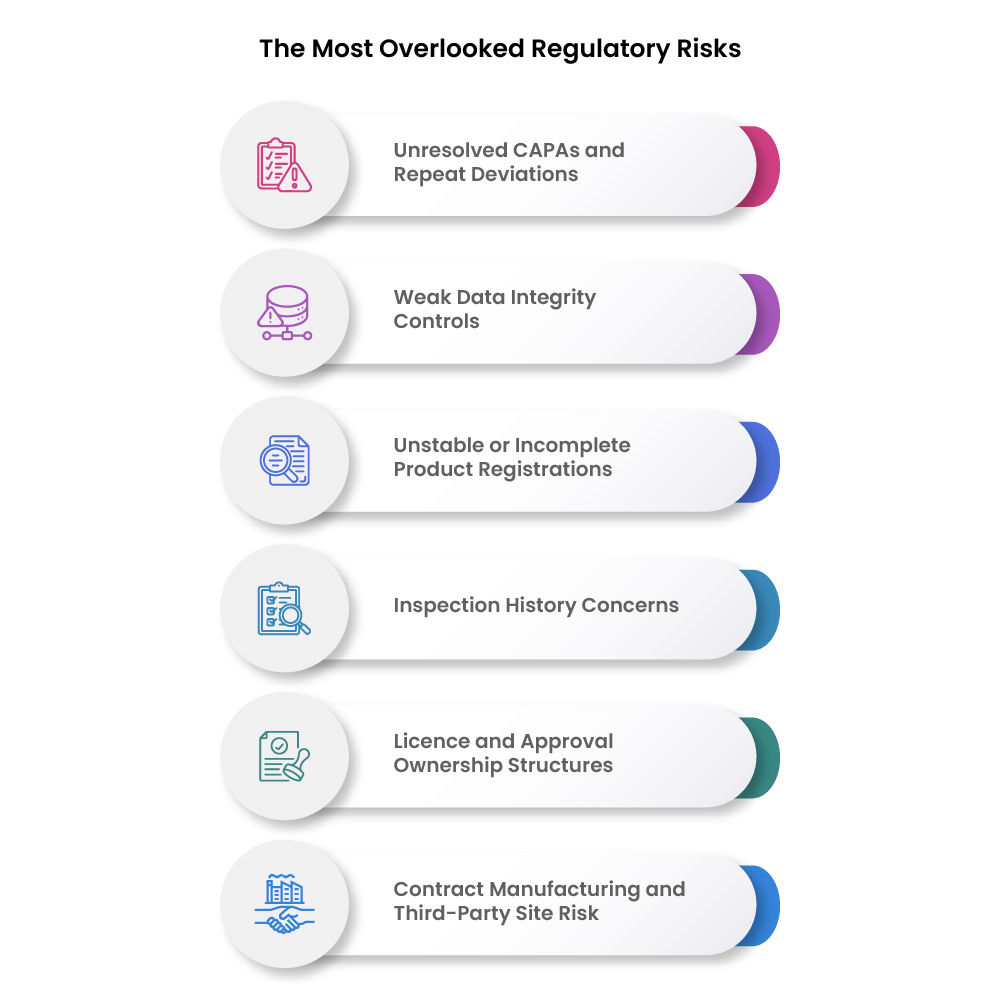
2. The Most Overlooked Regulatory Risks
Unresolved CAPAs and Repeat Deviations: A pattern of repeat deviations in the same area across multiple batches reveals a QMS that has not genuinely addressed root causes. Experienced reviewers always examine deviation trending data, not just individual CAPA records.
Weak Data Integrity Controls: A manufacturer may have no active falsification — but if audit trails are not routinely reviewed, access controls are inadequate, or electronic records do not meet ALCOA+ principles, the risk exposure is material. Data integrity remediation is expensive and subject to regulatory scepticism even after completion.
Unstable or Incomplete Product Registrations: Registrations may have lapsed, variations not filed for implemented changes, or dossier data not reflecting current manufacturing practice. These gaps create immediate post-acquisition remediation obligations.
Inspection History Concerns: A pattern of repeat observations across inspection cycles in the same area is a significant red flag. The quality of the manufacturer’s regulatory response letters is equally important — it reveals quality culture and management competence.
Licence and Approval Ownership Structures: In India’s Pharmaceutical sector, key manufacturing licences or product approvals are sometimes held in the name of an individual founder rather than the company entity. This creates change-of-control risk that can render approvals non-transferable without regulatory reapplication.
Contract Manufacturing and Third-Party Site Risk: A CMO inspection failure can result in product unavailability across the entire portfolio of clients operating from that site, regardless of the acquirer’s own compliance maturity.
3. How Compliance Maturity Links to Enterprise Valuation
- Major GMP remediation programmes consistently require USD 10–50 million or more for mid-to-large facilities. This erodes deal value materially when discovered post-close.
- Acquisition financing risk assessments increasingly require regulatory risk opinions as part of credit analysis.
- ESG frameworks applied to Pharmaceutical investments incorporate quality culture and compliance track record as non-financial performance indicators.
- Post-acquisition integration timelines are directly affected by compliance status. A mature target integrates in 12–18 months; a non-compliant one may require 3–5 years of intensive remediation.
4. How Manufacturers Can Improve Acquisition Readiness
- Commission a Pre-Transaction Regulatory Self-Assessment from the perspective of a sophisticated acquirer.
- Resolve Open CAPAs: Close all CAPAs with documented effectiveness checks; conduct formal deviation trend analysis for the preceding three years.
- Audit Your Product Registration Portfolio: Ensure all registrations are current, accurate, and held at the correct legal entity level.
- Strengthen Data Integrity Infrastructure: Conduct a formal DI audit across all GMP-critical systems; retain records of the remediation programme as evidence of proactive management.
- Build a Regulatory Inspection Track Record: A positive inspection history is a tangible asset in M&A.
- Engage Qualified Regulatory Advisory Support: Benchmark compliance posture against industry best practice before entering a transaction process.
Conclusion
Regulatory due diligence is a central determinant of Pharmaceutical deal value and post-acquisition success. Manufacturers who invest proactively in compliance maturity command premium valuations and deliver on post-transaction integration commitments.
CliniExperts provides pre-transaction compliance assessments, regulatory due diligence reviews, and post-acquisition remediation programmes for both manufacturers and investors.
Recent Posts
Schedule M Revisions: How Will They Impact Pharmaceutical Manufacturers?
What a Revised Schedule M raises GMP standards in India, what has changed for manufacturers, and how large firms and MSMEs can prepare for Compliance and CDSCO inspections. Revised Schedule M is the b..
CDSCO vs US FDA vs EMA: Why the Same Drug Faces Different Approval Requirements
A practical comparison of CDSCO vs FDA vs EMA, explaining how each regulator’s Drug approval process differs and why one Drug needs separate approvals in India, the US and Europe. A Drug approve..
How to Manufacture Class A & B Medical Devices in India: State FDA License Guide
A quick guide on the Class A & B Medical Device manufacturing license — covering State FDA rules, Form MD-3/MD-5, fees, and timelines for manufacturers. Every manufacturer entering India’s..
Get In Touch With Our Team
-
Email: contact@cliniexperts.com
-
Timings: 9 am to 6 pm (Monday to Friday)
-
Location: India: Delhi, Bangalore Global: USA, Singapore